Monday, 16 January 2023

Drug Master File

 Master records are authorized documents that contain the information to perform and control process operations and activities related to the manufacture of a product.

This article summarizes the requirements requested by the Food and Drug Administration (FDA) in its draft published in October for the master drug records or documents of pharmacological substances, packaging material, etc. The second section discusses the regulatory requirements for master documents submitted in sanitary registration applications in USA.

Guidance proposal for the industry

The FDA 510k Certification published in October the draft of the guide for the industry on the Master Drug File the guide provides the way to prepare the documentation on the facilities, processes or materials used in the manufacture, the manufacturing process, packaging and storage of medicines for human use.

The information described in the guide is not mandatory by any legal provision and the sending of the same is left to the consideration of the holders of the DMF. They can be used to support New Drug Applications (NDAs), Abbreviated New Drug Applications (ANDAs), and Investigational Drug Applications (INDs).

The guide proposes four types [1] of DMF's:

  • Type II. Pharmacological substance, intermediate and its raw materials for its manufacture.
  • Type III. Packaging materials.
  • Type IV. Excipients, colorings, flavorings, essence or material used for its preparation.
  • Type V. Accepted reference information.

Filing of the DMF is subject to the electronic filing requirements set forth in the Federal Food, Drug, and Cosmetic Act (FD&C Act) section 745A guidance and complying with the electronic common technical document (eCTD) format and Guidance ICH M4.

The DMF can be presented in original in the eCTD format, US FDA Registration for this the holders must have a pre-assigned number and a letter of presentation of technical administrative information based on the eCTD.

Tuesday, 27 December 2022

Clinical Evaluation for Medical Devices

 Clinical evaluation: The clinical evaluation report (CER) complies with the MEDDEV Guideline 2.7/1 rev. 4.

With the full implementation of the strict European Regulation (MDR 2017/745) in 2020, the issue of clinical evaluation has become a sore point for Medical Device manufacturers. Regardless of their risk class, all Medical Devices marketed in Europe are subject to the clinical evaluation process, in accordance with point 6 bis of Annex I of Directive 2007/47/CE. The Clinical Evaluation report (CER) alone is no longer enough.

While once considered just an element of the technical file, today the REC according to the latest version of the MEDDEV Guideline 2.7/1 (rev.4) is, in fact, the final phase of a carefully planned clinical evaluation process. Even if products are already CE marked, manufacturers must keep up with ever-higher standards because Notified Body (NO) control of clinical assessment and ERC will only intensify as 2020 approaches.

The MEDDEV Guideline 2.7/1 rev.4 defines clinical evaluation as a procedure that involves the collection, estimation, analysis and evaluation of the clinical data available to support the compliance of the Medical Device with the Essential Requirements (ER). In other words, the clinical data serve as evidence that, if used in accordance with the manufacturer's IFUs, the Medical Device is safe, has the claimed performance and any unwanted side effects, compared to the clinical benefit of the device, is considered acceptable.

So how can manufacturers ensure that their clinical evaluation process and RECs meet the latest regulatory expectations expressed in MEDDEV 2.7/1 rev.4?

Here are some helpful ideas to consider:

Define clear objectives in the CER. Identify relevant ERs supported by clinical trials (based on device specifications) and define the scope of clinical evaluation, i.e., specific and measurable goals related to safety, performance, and risk/benefit endpoints;

Define if a clinical investigation is required . Make sure that the available clinical trials are sufficient and adequate to answer all relevant ERs. As a general rule, high-risk and Class III devices should always undergo clinical investigation, as well as those based on innovative or unproven technologies, and those claiming a new clinical use;

Prepare a “bulletproof” Equivalence Proof . Based on the comparison between the device in question and several CE-marked equivalent devices available on the market, the demonstration of equivalence is established in terms of clinical, technological and biological parameters (defined by the manufacturer) and must be very detailed. at CER.

To provide a proper proof of equivalence, carefully documented information is required for each equivalent device. In addition, the differences between the device in question and the equivalent devices must be identified, disseminated, evaluated and justified;

Monitor post-marketing activities (PMS and PMCF). Plan an appropriate PMCF, document all findings in the CER, and ensure you have processes for PMS and PMCF in place. These must be "living" documents because the security and performance of a device must be confirmed throughout its life cycle;

Update the CER frequently . Define the CER update frequency and justify it. The CER is also a "living" document and therefore whenever new post-marketing surveillance data emerge that could influence the CER findings, they should be revised. Also, changes to the state of the art and current knowledge about equivalent devices should trigger a CER review. Devices used for comparison should always be kept relevant.

Under the strict requirements of the MDR that reflect the high expectations of the European Commission, NOs will question both the quality of clinical evidence (hence the criteria that include such data in clinical evaluation) and the basis for establishing equivalence with another device. . The stakes are high: they must ensure that only high-quality, safe and effective Medical Devices are placed on the European market.

Manufacturers preparing for new legislation have MEDDEV 2.7/1 rev.4 at their disposal to use as a roadmap to guide their transition. CERs based solely on demonstrating clinical equivalence will no longer suffice, especially for high-risk devices, which likely require clinical investigations. Only by aligning and integrating the CER with risk management, surveillance and post-market supervision activities will compliance with the requirements of the European MDR be achieved.

Monday, 21 November 2022

Responsibilities of the EU Authorized Representative

A European Authorized EU Representative is defined as a natural or legal person within the European Union who has accepted a written mandate from a manufacturer, located outside the EU, to act on its behalf in relation to specific tasks under MDR or IVDR. Specifically, Authorized Representatives play a critical role in ensuring device compliance and serving as a point of contact for European authorities and notified bodies.

MDR and IVDR describe the tasks that the manufacturer can delegate to the Authorized Representative and the conditions under which this can be carried out. This relationship must be covered by a precise mandate agreed between the EC REP and the manufacturer. The Authorized Representative will carry out the tasks specified in the mandate. A copy of the mandate will be provided by the Authorized Representative to the competent authority, upon request.

The Authorized Representative is responsible for defective medical devices along with the manufacturer, as well as any default by the manufacturer.

Mandatory tasks of the Authorized Representative according to MDR and IVDR

MDR and IVDR clearly describe the core role of the Authorized Representative and their obligations, which should be mentioned in the mandate:

  • Verify that the EU declaration of conformity and the technical documentation of the product have been drawn up and, where appropriate, that the manufacturer has followed the relevant conformity assessment procedure.
  • Keep a copy of the technical documentation, the EU declaration of conformity and, where appropriate, a copy of the corresponding certificate, including its modifications and supplements, available to the competent authorities.
  • Comply with FDA Registration obligations.
  • In response to the request of a competent authority, provide it with all the information and documentation necessary to demonstrate the conformity of the product.
  • Transmit to the manufacturer any request for samples or access to a product by a competent authority of the Member State in which the authorized representative has its registered office, and verify that said authority receives them or that access is granted.
  • Cooperate with the competent authorities in any preventive or corrective action taken to eliminate or, if this is not possible, mitigate the risks posed by the products.
  • Immediately notify the manufacturer of claims and reports from health professionals, patients or users about alleged incidents related to a product for which it has been designated.

Thursday, 20 October 2022

EU Representative For Medical Devices – I3CGLOBAL

Due to the increasingly common presence of imported medical devices in the European Union, the figure of the Authorized Representative (EU Representative) is increasingly common.

This figure must guarantee compliance with current regulations on medical devices for medical devices manufactured outside the European Union. Regulation 2017/745 on medical devices establishes the obligation for all manufacturers of medical devices established outside the European Union to have a permanent authorized representative who serves as an interlocutor for the European health authorities and has a documentary file with technical information of medical devices (Medical Device TechnicalFile among others).

 The authorized representative must be a natural or legal person based in a member state of the EU and appear on the labels of medical devices with CE marking for marketing in the EU. 

The appointment of this EC REP must be in writing together with the acceptance of it and in the case of class I products, this appointment must be communicated to the competent authority in the field of medical devices so that said authority may register it in the Register of Responsible Parties. marketing of sanitary products.

The authorized representative designated by the manufacturer of medical devices must keep the documentation regarding the products for 5 years from the cessation of marketing of the medical device and 15 years in the case of implantable medical devices (intraocular lenses, prostheses of breast, hip, joints, spine, etc.).

 I3CGLOBAL has extensive experience in consulting for medical devices and especially in international trade, acting for its clients who manufacture medical devices based outside the EU as a permanent European Authorized Representative (EC REP) in the EU.

 Why Choose I3CGLOBAL for European Authorized Representative?

 We are the Best, Fastest, and Most Cost-Effective Medical Device EU Representative in Europe for EU FDA Registration and Free Sale Certification

 GET IN TOUCH


Friday, 23 September 2022

Do I Need an FDA Number? The Answer May Surprise You:

 A blog about needing an FDA registration number and how you can get one.

Are you a food manufacturer? Do you make food products, treat surfaces or provide packaging material that comes in contact with food? Then you will definitely need an FDA registration number. Learn more about getting an FDA number.

There are a lot of questions that need to be answered when it comes to getting your product on the market.

One of these questions is whether or not you need an FDA registration number. Many people are under the impression that if their product is sold online, then they don't need to worry about getting one.

But that couldn't be further from the truth. If you sell products online, there are some things you should know about getting an FDA registration number and why it's so important for your business.

What Do I Need to Know About Getting An FDA Registration Number?Clinical Evaluation

It's important to note that the FDA does not require all businesses to have a registration number, only those who manufacture or process food in some way.

This means that if your business is selling food products, you will most likely need one in order to legally sell them in any state within the United States.

However, this doesn't mean that all sellers need one—just those who make and sell their own items rather than purchasing them from another company and reselling them through their website or store front location (such as Walmart).

However, if you're selling your products nationally or internationally, it's important to have an FDA registration number.

If you're selling directly to consumers, the FDA registration number is not required. However, if you're selling your products nationally or internationally, it's important to have an FDA registration number.

In addition to bringing credibility to your business, having an FDA number allows you to sell your products over state lines and into other countries without having to complete a new application for each state.

The Food and Drug Administration (FDA) is a federal agency that regulates food safety, dietary supplements and medical devices in the United States. The FDA registers businesses that manufacture or distribute foods, drugs, cosmetics and other consumer products like dietary supplements.

The FDA registration process is quick and easy and anyone can do it.

The FDA requires that all companies who sell cosmetics in the US register their business with the agency. However, there are some exceptions to this requirement. If you're just selling your products at local craft fairs or festivals, you probably don't need a registration number. If you're selling online only (and not in person), you may also be exempt from this requirement.

If you want to sell your cosmetics through brick-and-mortar stores or through websites other than yours (for example, on Amazon), then yes, you'll need an FDA registration number. If you have any question regarding US FDA Registration process you can simply visit our website www.i3cglobal.com

Monday, 22 August 2022

BIOCOMPATIBILITY TEST FOR MEDICAL DEVICE

 Biocompatibility is a measurement of how compatible a device or material is with a biological system. The ISO 10993-1: 2018 standard defines biocompatibility as the ability of a medical device or material to perform with an appropriate host response in a specific application".

The term "biocompatibility" describes how a medical device interacts with a patient's tissues and physiological systems while being used to treat them. One aspect of a device's overall safety evaluation is an assessment of biocompatibility. Analytical chemistry, in vitro tests, and animal models are used to investigate the biocompatibility of devices. A device's biocompatibility is influenced by a number of factors, including:

  • the physical and chemical composition of its constituent parts
  • the time span of that exposure
  • patient tissue types that will be exposed to the device

The purpose of performing biocompatibility testing is to determine the fitness of a device for human use and to see whether the use of the device can have any potentially harmful physiological effects. The goal of device designers is to minimize risk while maximizing benefit to patients.

Biocompatibility testing can be divided into three steps: planning, conducting the test, and analyzing the results. Those three steps to demonstrate biocompatibility are stated below:

  1. Create a Biological Evaluation Plan (BEP). Data on the materials used to make the device should be gathered early in the planning process. The type of testing that is necessary will be determined by a biological evaluation plan (BEP). This examines your device and its materials, identifies potential risks, and suggests possible evaluations or testing to address those risks. The BEP can serve as your initial risk assessment, and you can also share it with the FDA during a free pre-submission discussion. 
  1. Evaluation and testing of devices. This entails performing the tests identified in your BEP. These are typically a combination of in vivo or in vitro biological tests, chemistry tests, and toxicological risk assessments, as well as written assessments based on scientific literature.
  1. Create a Biological Assessment Report (BER). The results of all tests and evaluations are summarized here. This, along with the test results, is submitted to the manufacturer.

  • APPLICABLE STANDARD -

As stated by the International Organization of Standards (ISO 10993-1:2018), “The primary aim of this part of ISO 10993 is the protection of humans from potential risks arising from the use of medical devices. Before performing biocompatibility testing, it is important for the manufacturer to gain a better understanding of the device materials, device manufacturing, sterilization, and other processes. The ISO 10993-1:2018 standard emphasizes chemical characterization before proceeding to in vitro and in vivo biocompatibility testing.

EN ISO 10993-1:2020 - Biological evaluation of medical devices - Part 1: Evaluation and testing within a risk management process.

If a medical device does not have direct or indirect contact with the patient, then ISO 10993-1 does not apply.

ISO 10993 consists of the following parts, under the general title biological evaluation of medical devices:

⎯ Part 1: Evaluation and testing within a risk management process

⎯ Part 2: Animal welfare requirements

⎯ Part 3: Tests for genotoxicity, carcinogenicity, and reproductive toxicity

⎯ Part 4: Selection of tests for interactions with blood

⎯ Part 5: Tests for in vitro cytotoxicity

⎯ Part 6: Tests for local effects after implantation

⎯ Part 7: Ethylene oxide sterilization residuals

⎯ Part 9:Framework for identification and quantification of potential       degradation products

⎯ Part 10: Tests for irritation and skin sensitization

⎯ Part 11: Tests for systemic toxicity

⎯ Part 12: Sample preparation and reference materials

⎯ Part 13: Identification and quantification of degradation products from polymeric medical devices

⎯ Part 14: Identification and quantification of degradation products from ceramics

Read More:- FDA 510k

Wednesday, 20 July 2022

FDA Drug Establishment Registration

 Drug Establishment Registration is a program that helps ensure the safety and effectiveness of drugs by requiring manufacturers, packagers, and distributors to register their establishments with the FDA. The registration process requires the submission of information about the manufacturing practices, quality systems, and facilities used at an establishment's place of business.

If you're an online retailer and would like to sell pharmaceutical products in the U.S., then you'll most likely have to become FDA Drug Establishment Registered. Once a drug is approved by the FDA, it is assigned a unique identifying code. Once registered, products can be sold in the U.S. without restrictions on advertising or distribution channels.
Drug Establishment Registration Process

Drug Establishment Registration is a process of registering and obtaining a license for the establishment where drugs are manufactured or processed.

This process is conducted by the Food and Drug Administration (FDA) of the Philippines. The purpose of this registration is to ensure that the drugs manufactured in the country are safe for human consumption and are made according to FDA standards.

The applicant must submit an application form along with other requirements such as:

Copy of valid business permit

Proof of tax payment

Proof that they have complied with all requirements of their trade association/association

Drug master file (DME), which contains information about the drug product, including its composition, quality control processes, and manufacturing specifications
Note: Manufacturers must register with the FDA with their own unique Drug Establishment Registration number (DER), which will be used on all labels and packaging for any products they produce. This includes certain types of packaging materials, as well as any ingredients or additives they use in their manufacturing process.

Importers must also register with the FDA, but they do not need to use a DER number on their packaging materials because they do not make the drugs themselves—they just import them from other countries where there may be fewer regulations to ensure quality control.

Every chemical component used as a medicine is considered as a drug under US FDA regulation. This could be used for local application or for oral consumption, or for intravenous usage. It could be for human use or animal use. FDA Drug Facility is any establishment that manufactures, repackages, or re-labels drug products in the United States or foreign location.

Who Need Drug Establishment Registration?

Any company that manufactures, repackages, or re-labels drug products in the United States must register with FDA. The company could be of domestic or foreign origin. Both domestic and foreign establishments must register with FDA. These drug products could be for animal use or human use.

Unless Exempted by law all drug manufactures (Like OTC Drug , Prescription Drugs, Generic Drugs, API, Compounding Drug, Sterilization Facilities, Testing Labs ) must complete FDA Establishment registration within 5 days of beginning of operations.

In Addition to Establishment Registration facilities are required to list their drugs with FDA and this listing needs to be renewed every year before 31st December.

How to Register for Drug Establishment?

The minimum information needed while registering will be:

  • Name of establishment(s) manufacturing or processing the listed drug and the type of operation(s) performed.
  • DEA schedule.
  • Route(s) of administration.
  • Inactive ingredients.
  • Marketing information
  • Information related to the application or OTC monograph citation number Package size and type.
  • Flavor, Color, Active Ingredients.

Electronic Drug Listing (EDL) in the FDA database will also need SPLs

The Structured Product Labelling (SPL) is a document markup standard approved by Health Level Seven (HL7) and adopted by FDA as a mechanism for exchanging product and facility information.
In addition to this, the application needs:

  • Unique Ingredient Identifiers (UNII)
  • Data Universal Numbering System (D-U-N-S®) Number
  • If all this jargon seems too much to handle, let’s make it easy.

Remember: Registering at a drug establishment is an essential step in the distribution of pharmaceutical products. It's an important piece of making sure your business complies with all federal and state laws. In this guide, we're going to discuss why it's so important that you register your establishment as well as what steps are involved in registering.

We are the leading experts in this field for 30 years. Let us handle the process, you can handle the profits.Share your contact information with us and we will revert you asap.

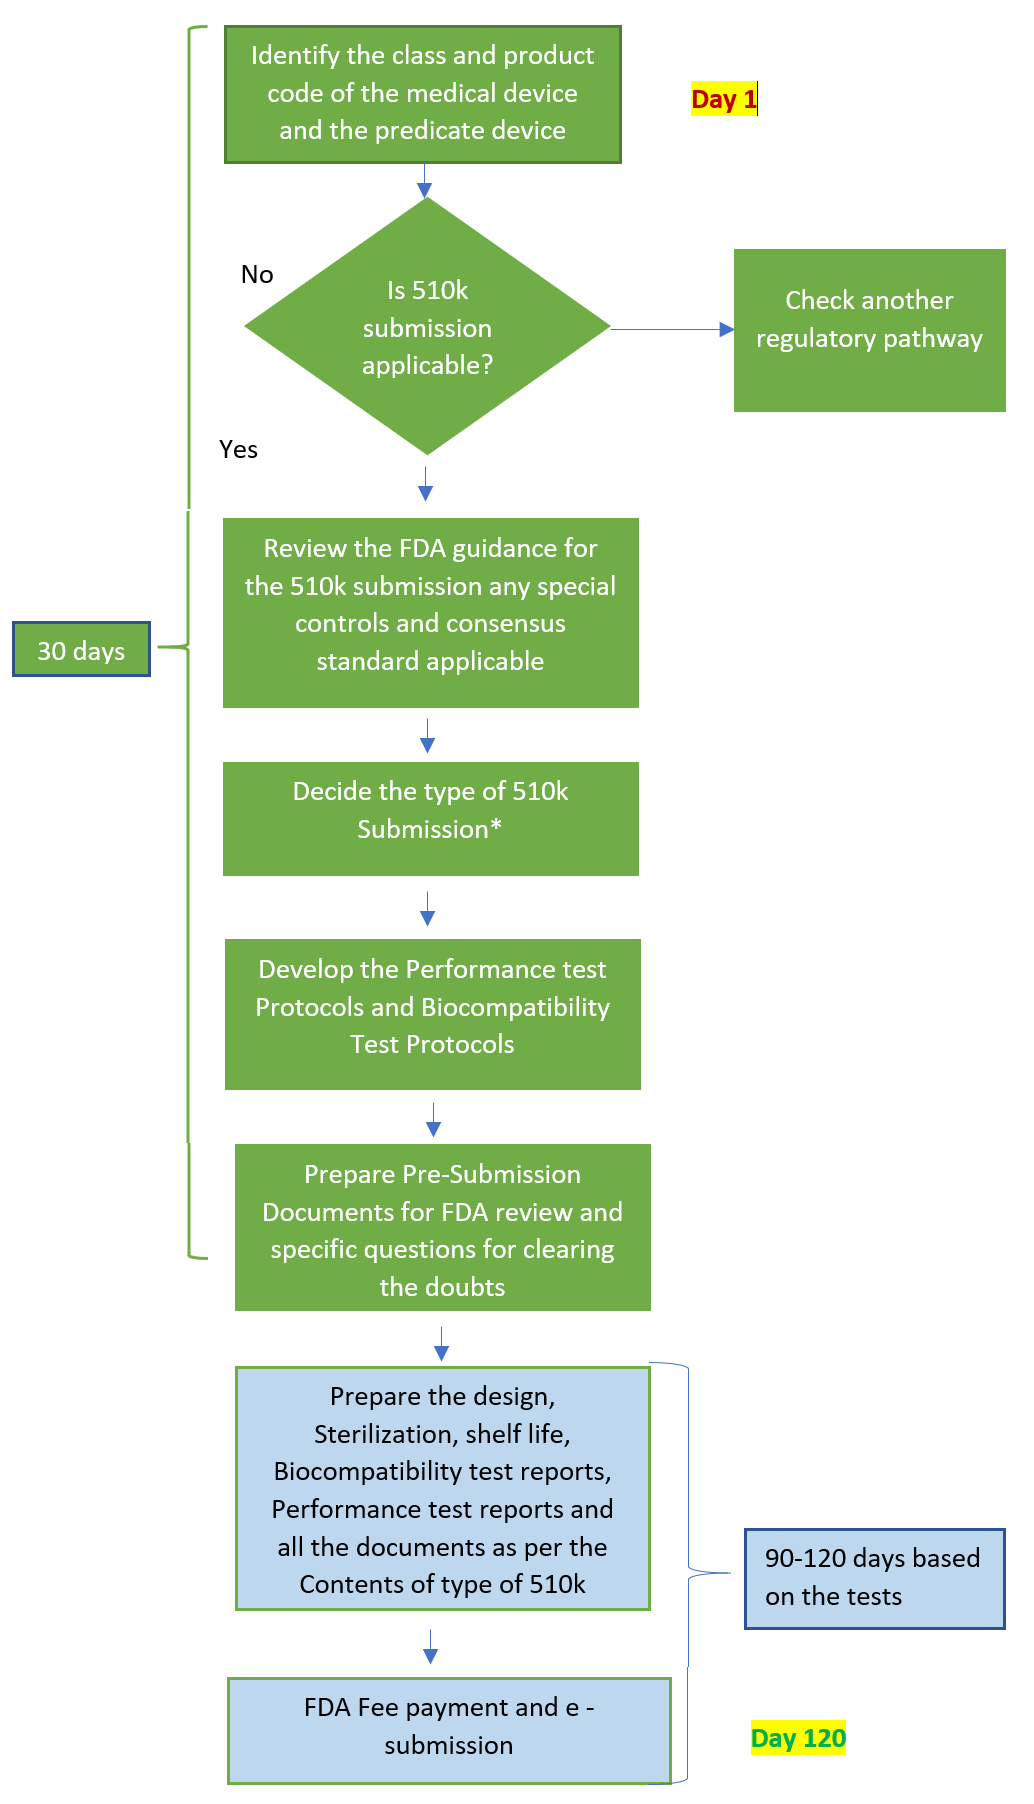
Find more details here at :- 510k submission